Chapters and reference
Module II · Single-sequence circular maps · Section 9 of 12
2.8 Restriction searches, cuts and labels
On this page
Searching cuts and choosing a display subset are separate steps. Compare leaders, text content, order and side on the same three pUC19 hits, without editing DNA or cut coordinates.
1 Retain cut coordinates and search assumptions
Run independently with explicit EcoRI, BamHI and HindIII and methylation=“none”. This is a search assumption, not a measurement of sample methylation. enzyme_site_count refers to the complete input sequence and survives display filtering.
R · Example code
finish_sites <- find_restriction_sites(finish_dna,
  enzymes = c("EcoRI", "BamHI", "HindIII"), methylation = "none")
R · Example code
finish_sites[c("enzyme", "position", "enzyme_site_count", "methylation_status")]| enzyme | position | enzyme_site_count | methylation_status |
|---|---|---|---|
| EcoRI | 396 | 1 | unknown |
| BamHI | 417 | 1 | unknown |
| HindIII | 447 | 1 | unknown |
2 Compare leaders at the same cut sites
Default radial uses text bounds and circular order, with radial stubs and fan connections where needed. Elbow uses one bend; straight connects the tick tip to the final label edge. Only leader changes across the three figures; cut sites and label grouping are fixed.
R · Example code
p_comparerestrictionradial <- finish_recognition_base +
  geom_restriction_site(data = finish_sites, leader = "radial")
view_ggchord(p_comparerestrictionradial, width = 9, height = 8, units = "in")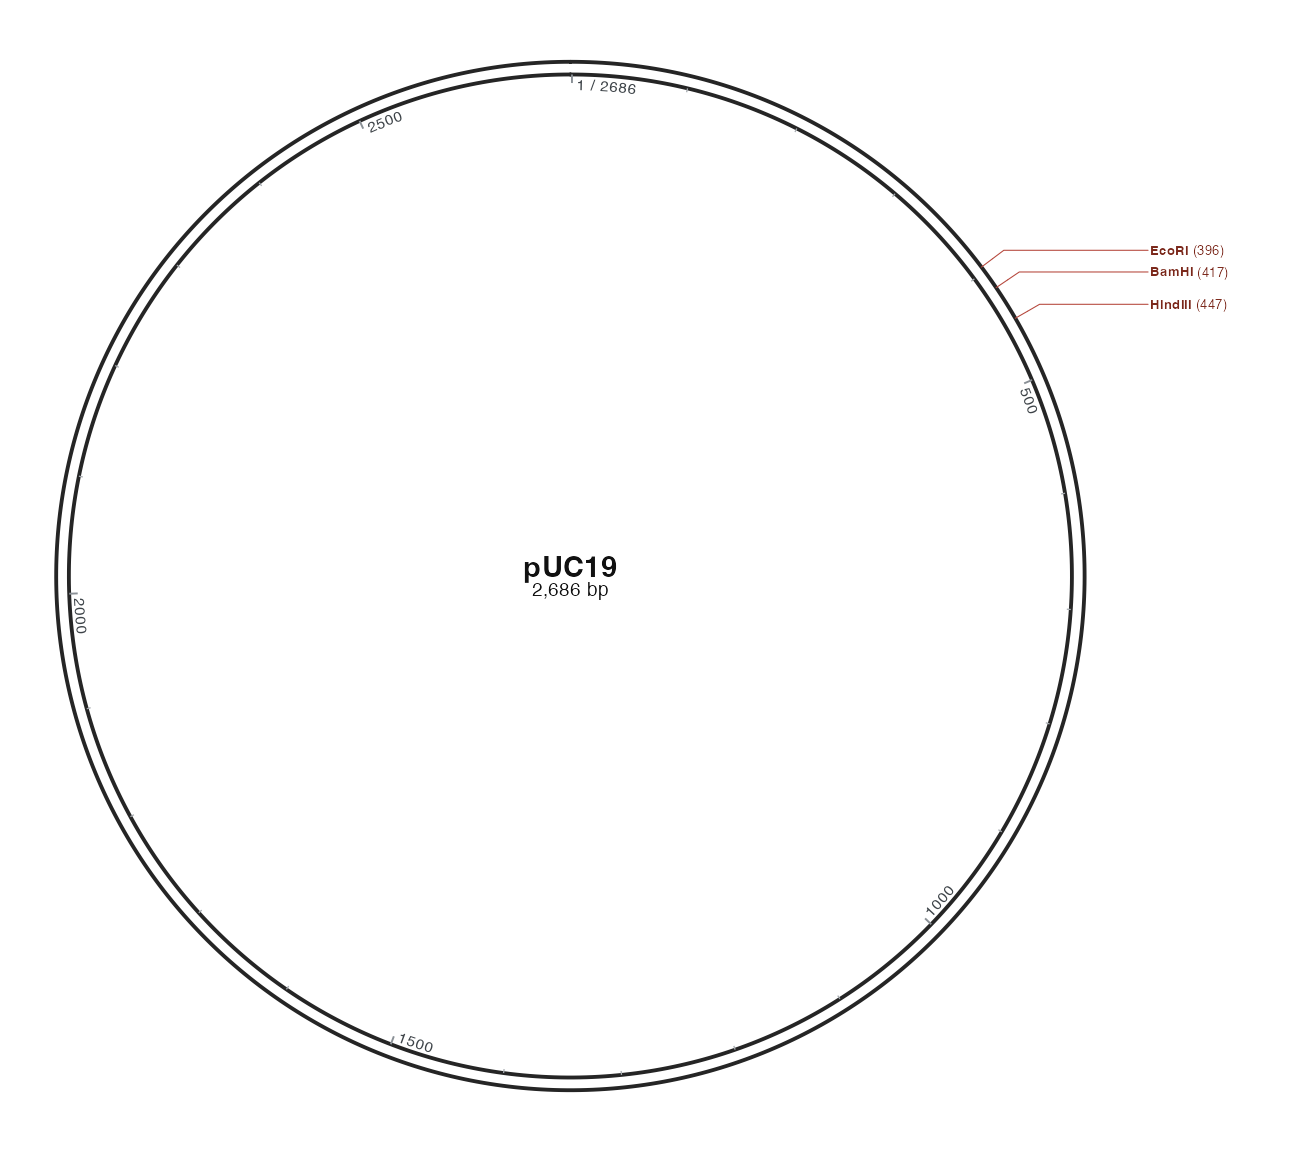
R · Example code
p_comparerestrictionelbow <- finish_recognition_base +
  geom_restriction_site(data = finish_sites, leader = "elbow")
view_ggchord(p_comparerestrictionelbow, width = 9, height = 8, units = "in")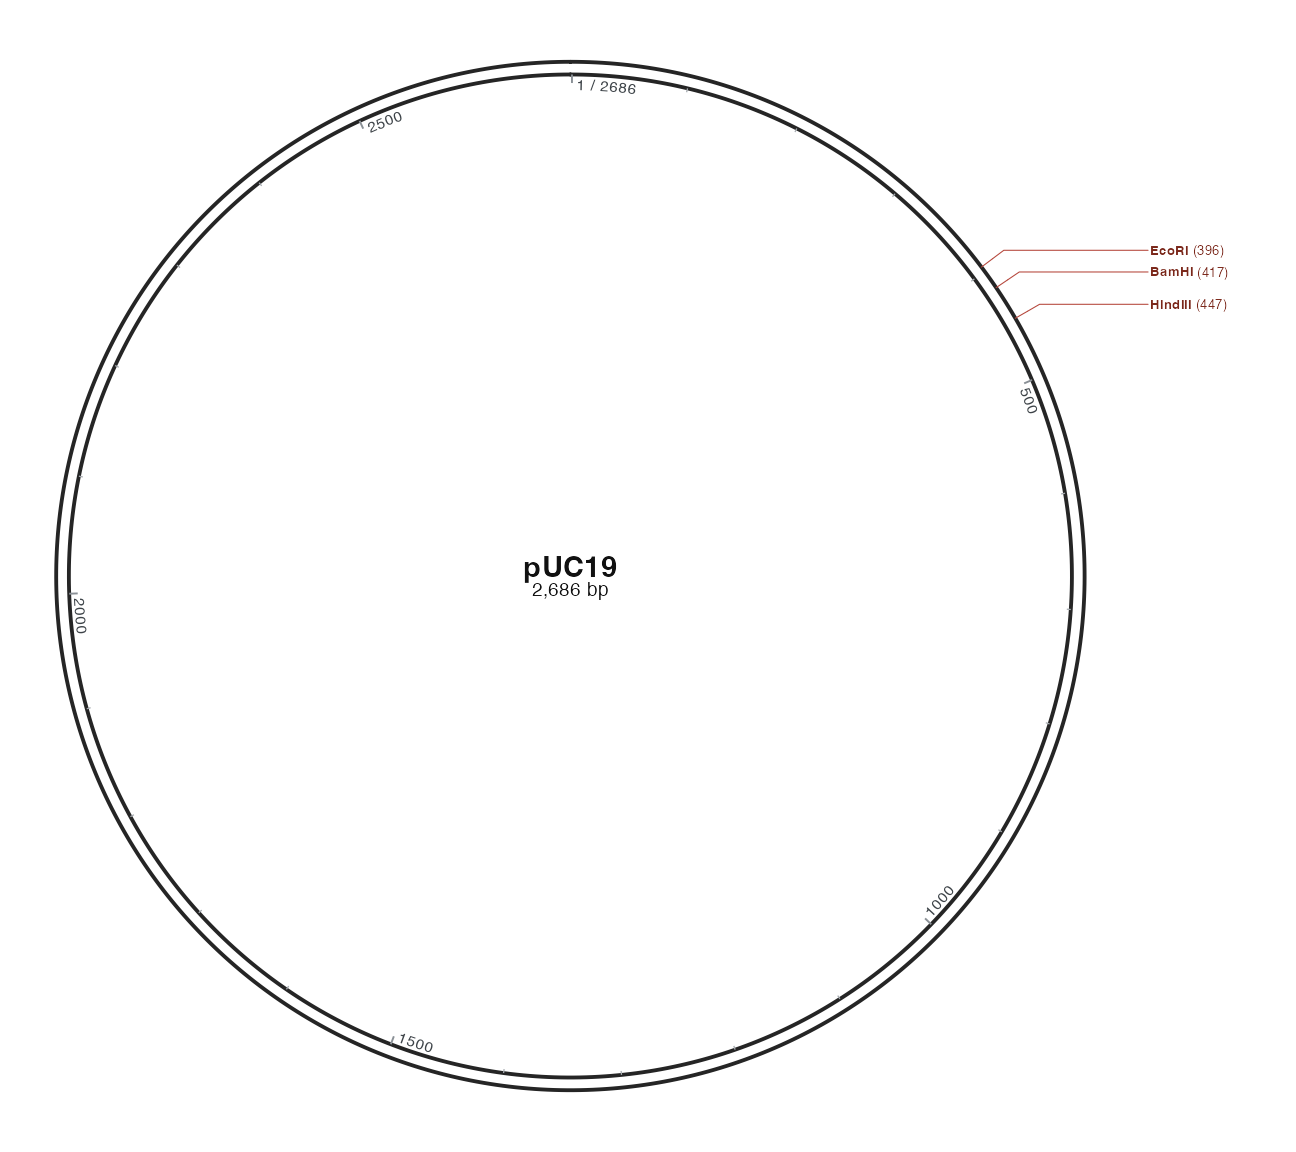
R · Example code
p_comparerestrictionstraight <- finish_recognition_base +
  geom_restriction_site(data = finish_sites, leader = "straight")
view_ggchord(p_comparerestrictionstraight, width = 9, height = 8, units = "in")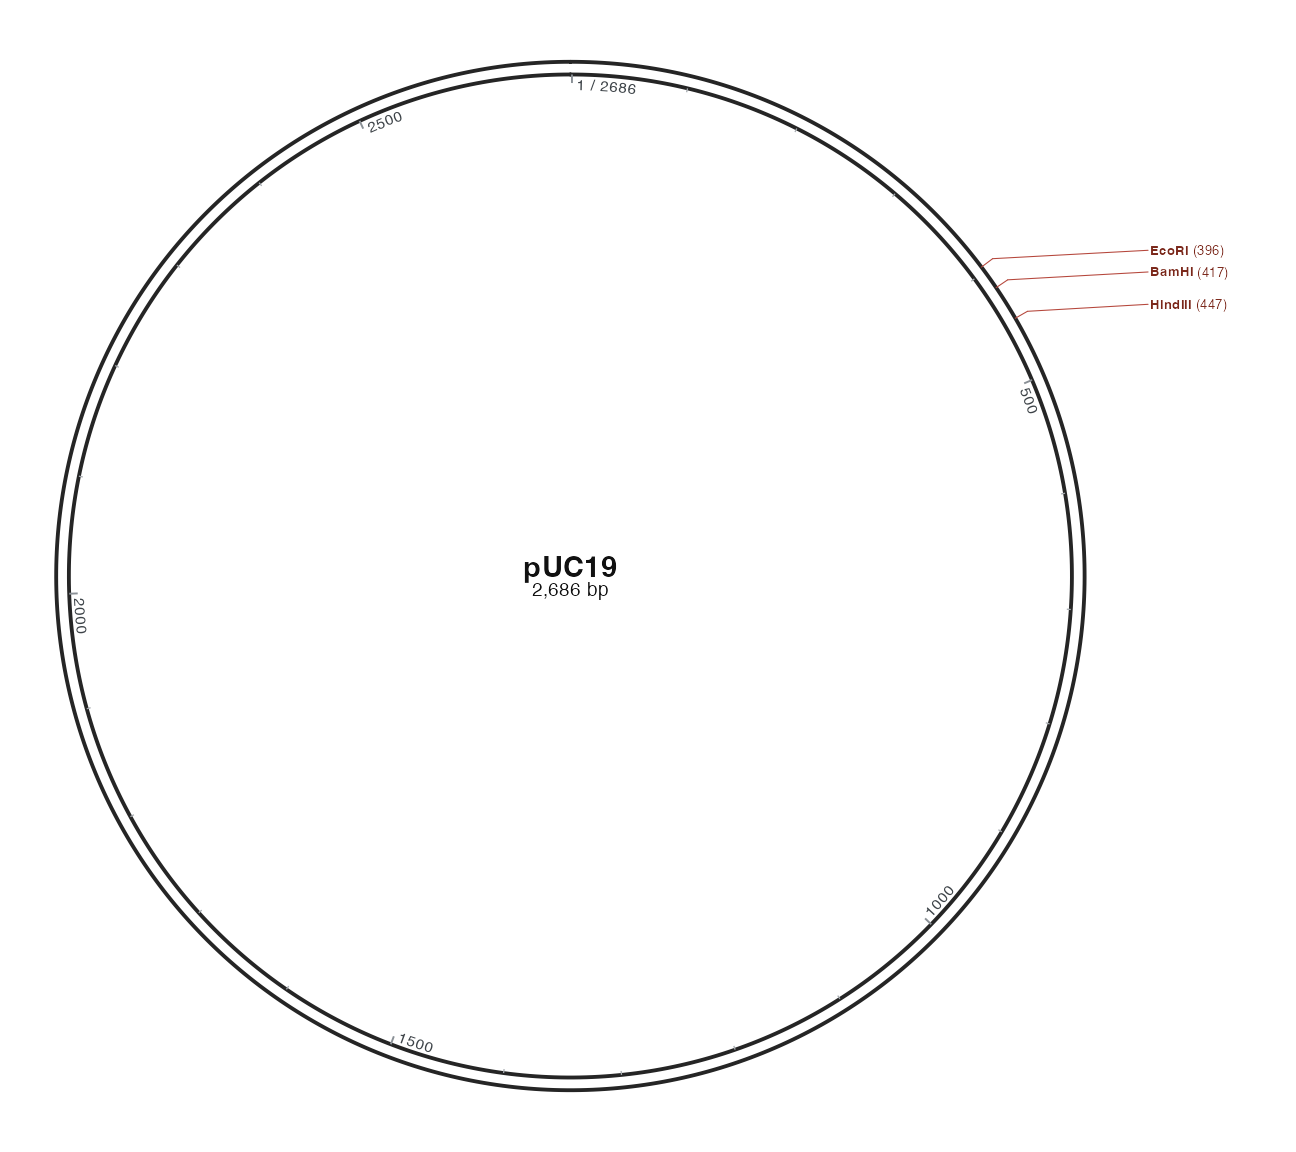
3 Change content, order and side separately
Each figure returns to the radial baseline, changing only label_style, label_order or label_side. Default bold enzyme names indicate one cut in the complete target; explicit fontface replaces that automatic distinction. Positions come from computed cleavage, not motif midpoints.
R · Example code
p_comparerestrictionenzyme <- finish_recognition_base +
  geom_restriction_site(data = finish_sites, leader = "radial", label_style = "enzyme")
view_ggchord(p_comparerestrictionenzyme, width = 9, height = 8, units = "in")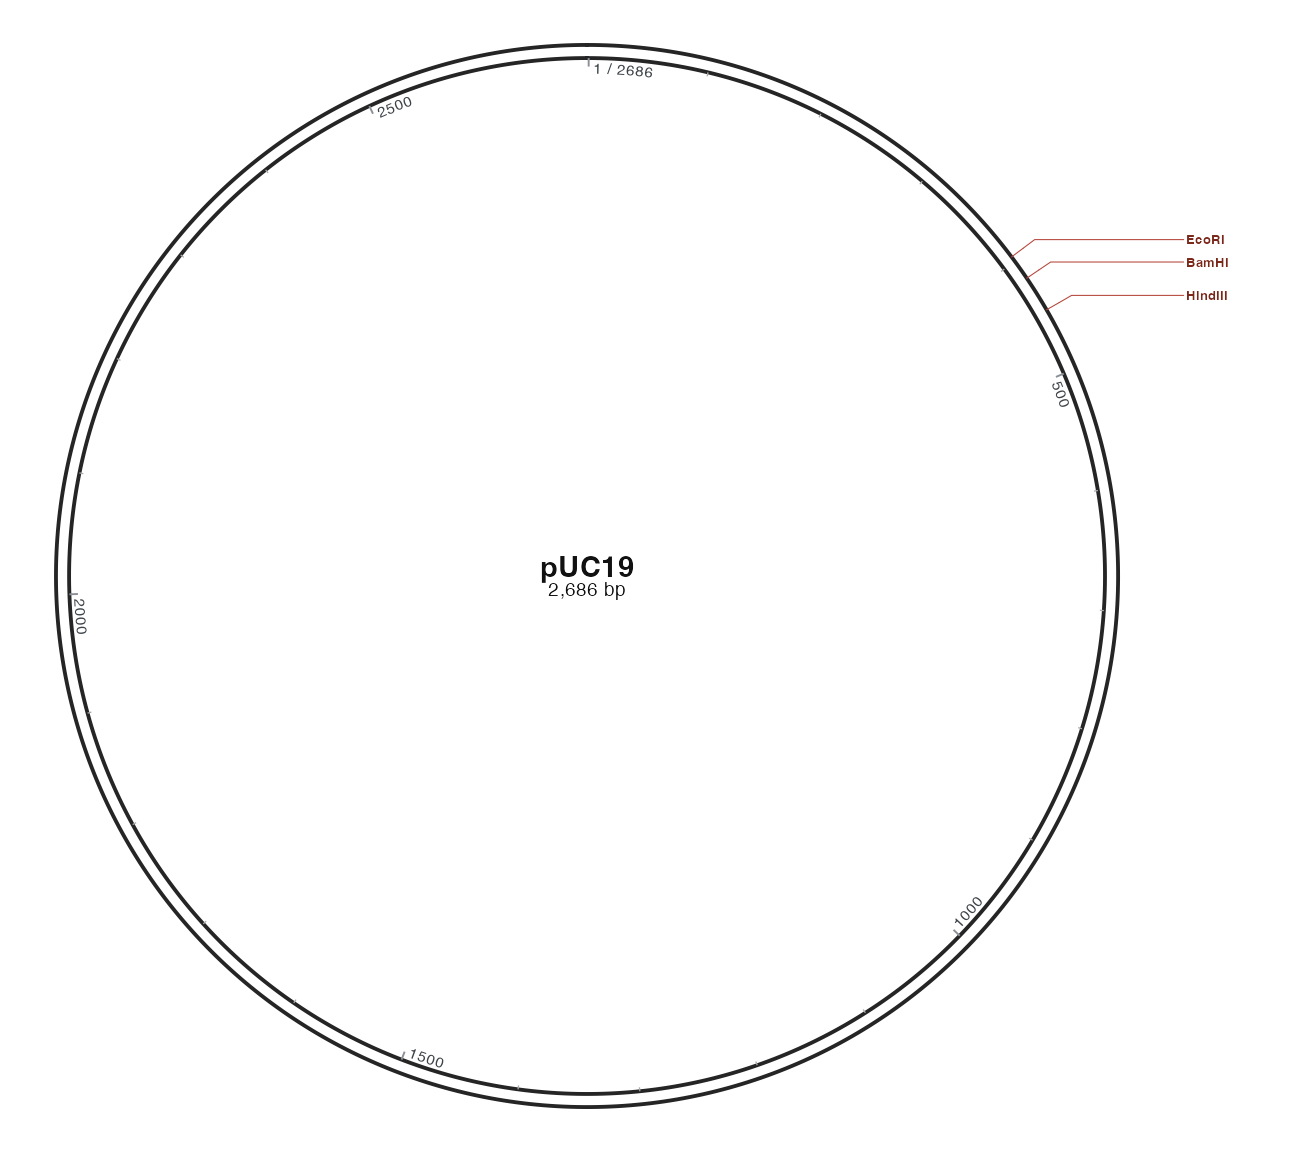
R · Example code
p_comparerestrictionposition <- finish_recognition_base +
  geom_restriction_site(data = finish_sites, leader = "radial", label_style = "position")
view_ggchord(p_comparerestrictionposition, width = 9, height = 8, units = "in")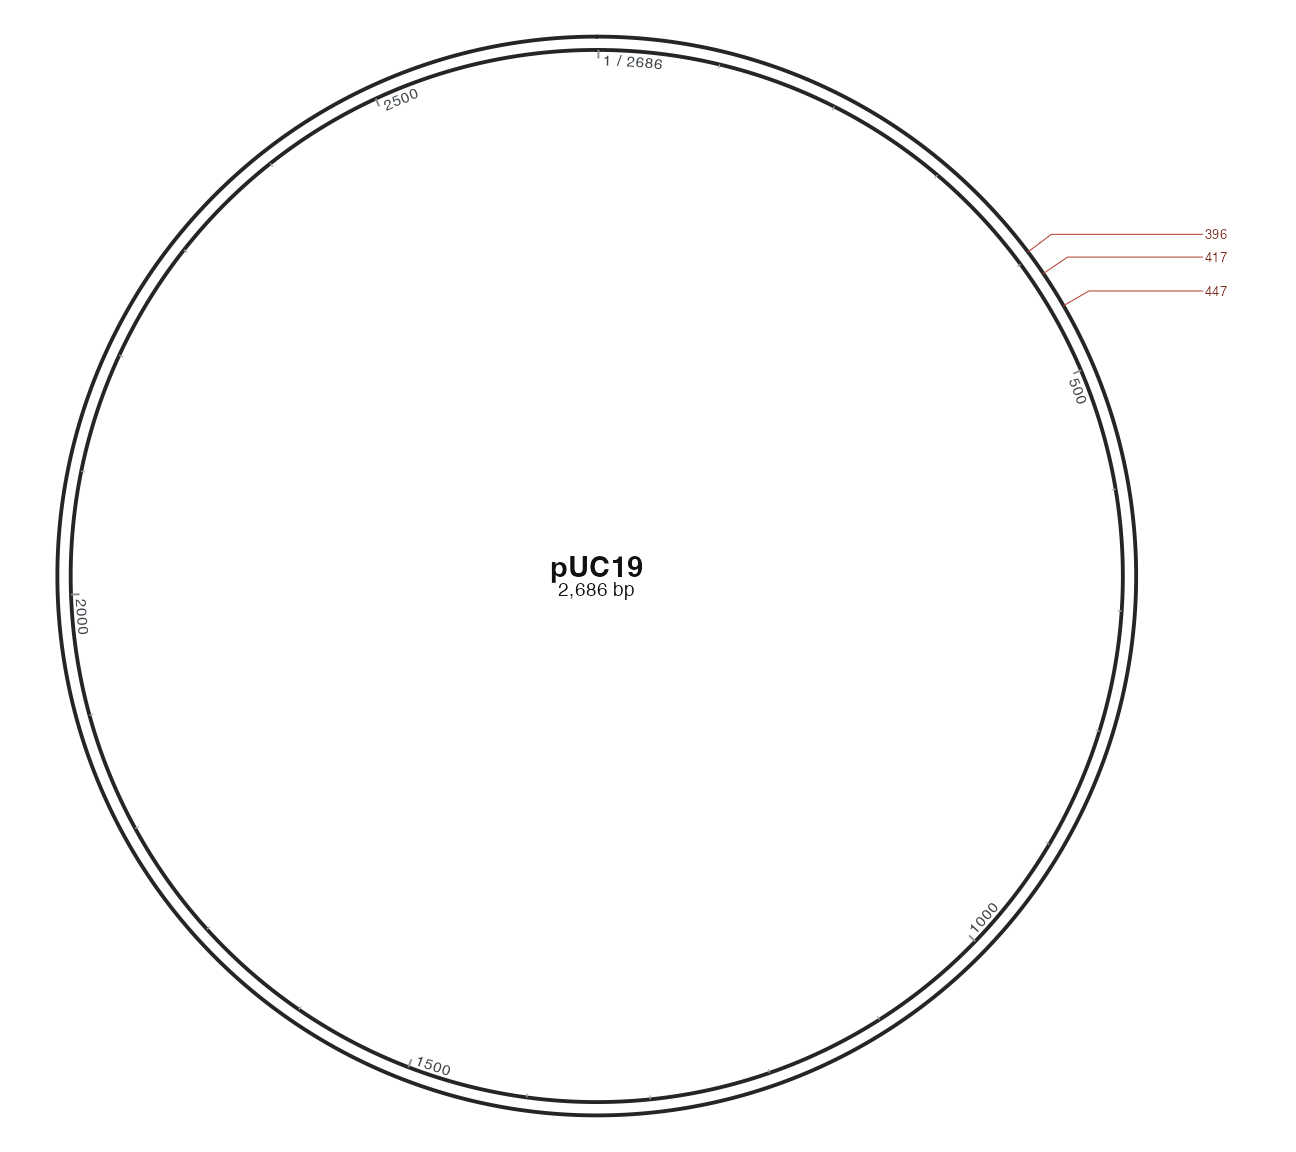
R · Example code
p_comparerestrictionorder <- finish_recognition_base +
  geom_restriction_site(data = finish_sites, leader = "radial", label_order = "position_enzyme")
view_ggchord(p_comparerestrictionorder, width = 9, height = 8, units = "in")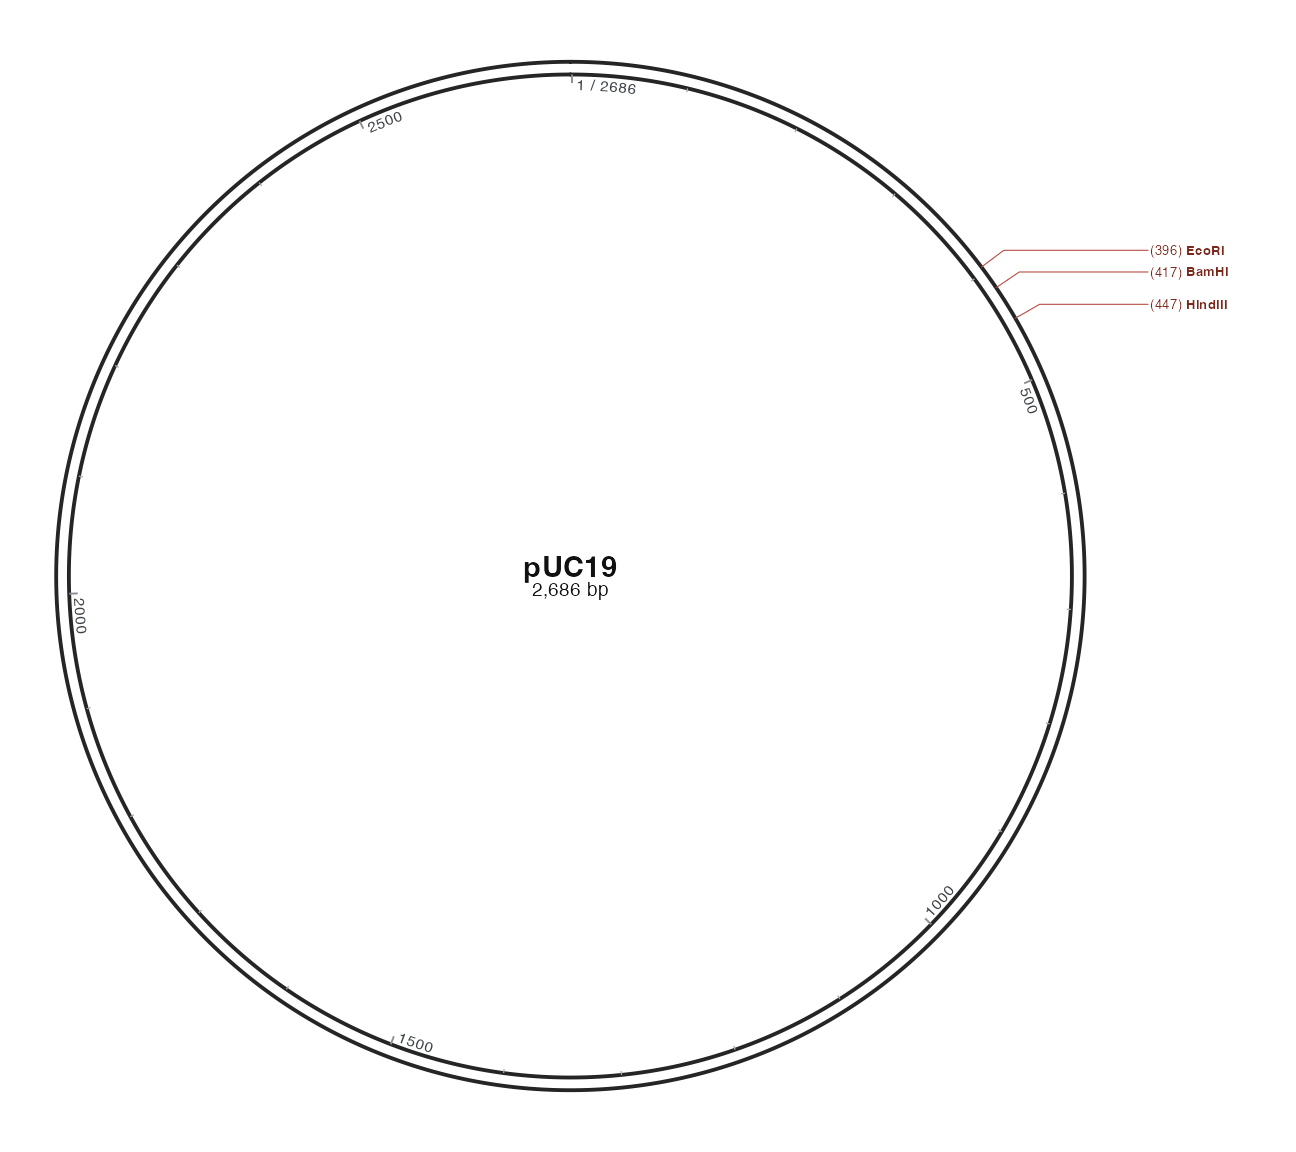
R · Example code
p_comparerestrictioninside <- finish_recognition_base +
  geom_restriction_site(data = finish_sites, leader = "radial", label_side = "inside")
view_ggchord(p_comparerestrictioninside, width = 9, height = 8, units = "in")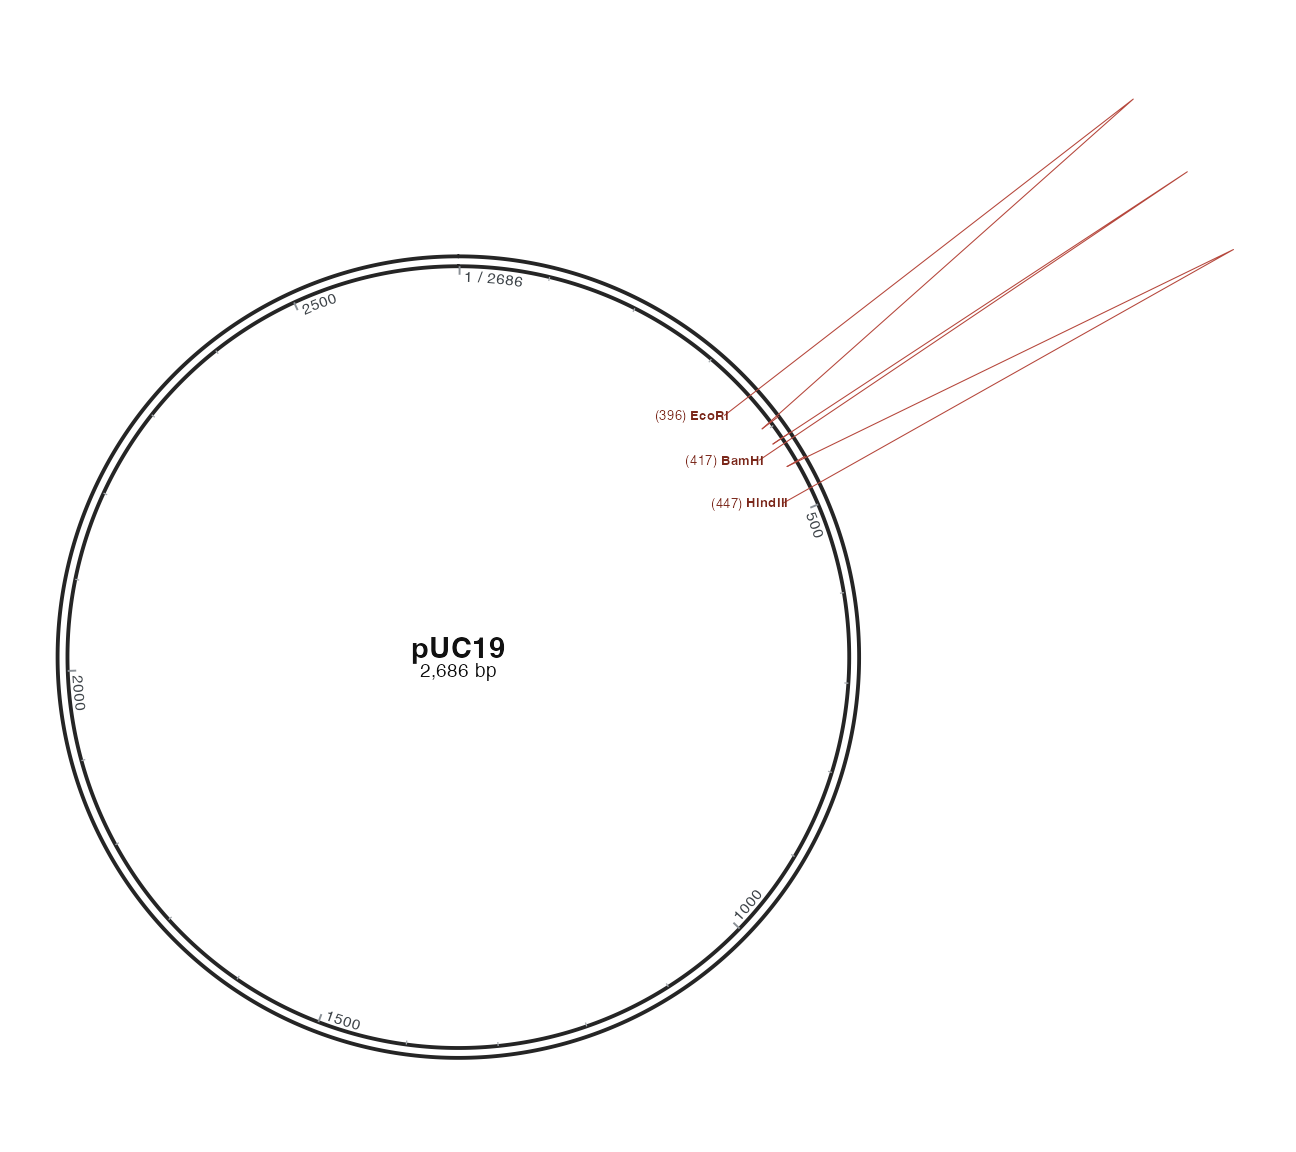
4 Key parameters and filtering meaning
| Parameter | Default | Effect |
|---|---|---|
| label_style | enzyme_position | enzyme_position/enzyme/position |
| label_order | auto | auto/enzyme_position/position_enzyme; automatic mirrored ordering |
| label_side | outside | outside/inside/auto |
| leader | radial | radial/elbow/straight; never changes cuts |
| min_label_gap | NULL | Device-derived default; explicit genomic-fraction spacing |
| tick_length | 0.04 | Local-normal tick length |
| label_offset | 0.18 | Local-normal text offset |
| linewidth / label_size | 0.28 / 3.2 | Stroke width versus text size |
| fontface / family | NULL / NULL | Explicit fontface overrides unique-cutter bolding |
find_restriction_sites returns matches and cuts; filter_restriction_sites selects a display set. unique, six_plus, unique_6plus and reference have different conditions. Reference uses saved sets only for exact reference targets. methylation=auto applies audited backgrounds only to exact built-in references; state assumptions for unknown samples.
5 Dense maps and usage limits
Increase only min_label_gap and inspect outward label/leader movement; then restore defaults and set label=FALSE for ticks only. Retain full search results separately from the display subset. Increase export dimensions instead of deleting source observations to fit a small image. Recheck crowding after combining features and primers in dense pETDuet-1. Computed cuts do not validate a digestion experiment.